14.4. PbTe (non-analytic correction)#
At a glance
- Goal:
Include the non-analytic correction in the phonon dispersion of the polar semiconductor PbTe.
- You will learn:
how to let alm build the supercell from the primitive cell with
SUPERCELLhow to create the displaced structures with
displace.pyfrom the primitivePOSCARhow to create the
BORNINFOfile (dielectric tensor and Born effective charges) from a DFPT calculationhow the
NONANALYTICoptions 1, 2, and 3 differ
- Prerequisites:
VASP (the VASP outputs are provided in
vasp_outputs/) andPyYAML(used bydisplace.py).- Example files:
example/PbTe
This part demonstrates how to include the non-analytic correction in phonon and thermal conductivity calculations of polar semiconductors.
All input files can be found at the example/PbTe subdirectory.
The VASP inputs are in vasp_inputs/, the VASP outputs of the force calculations in vasp_outputs/,
and the files generated in each step in reference/, so you can skip any step by copying the corresponding file.
Let’s move to the example directory
$ cd ${ALAMODE_ROOT}/example/PbTe
1. Build a supercell and generate displacements#
In this tutorial, we build a 4x4x4 supercell of the primitive cell of PbTe containing 2 atoms. The crystal structure of the primitive cell in the VASP POSCAR format is
PbTe
6.443068
    0.00000000000000    0.50000000000000    0.50000000000000
    0.50000000000000    0.00000000000000    0.50000000000000
    0.50000000000000    0.50000000000000    0.00000000000000
Pb Te
1 1
Direct
    0.00000000000000     0.00000000000000     0.00000000000000
    0.50000000000000     0.50000000000000     0.50000000000000
The lattice constant \(a = 6.4431\) Å was obtained by relaxing the primitive cell with the PBEsol functional
(INCAR.relax and KPOINTS.relax in vasp_inputs/; the relaxation was run twice, the second time starting from the CONTCAR of the first, which left a residual stress of 0.003 kB).
This step is optional here, since the relaxed POSCAR is provided, but for your own material, please relax the primitive cell first with the same DFT settings as the force calculations:
phonon frequencies of polar materials are sensitive to the lattice constant.
Copy the primitive cell to the working directory
$ cp vasp_inputs/POSCAR .
We do not need to build the supercell ourselves. alm builds it from the input structure with the SUPERCELL tag.
The input for alm (ALM0.in) is
&general
 PREFIX = pbte444
 MODE = suggest
 STRUCTURE_FILE = POSCAR
 SUPERCELL = 4
/

&interaction
 NORDER = 1
/

&cutoff
 *-* None
/
Here, STRUCTURE_FILE reads the lattice vectors, the elements, and the atomic positions from POSCAR,
so &cell and &position are not needed.
SUPERCELL = 4 means a 4x4x4 supercell of the input cell, \((\boldsymbol{a}_s, \boldsymbol{b}_s, \boldsymbol{c}_s) = 4(\boldsymbol{a}_p, \boldsymbol{b}_p, \boldsymbol{c}_p)\), which contains 128 atoms.
Since PRIMCELL is not given, it takes the default value (identity matrix), and the input cell itself is used as the primitive cell.
NORDER = 1 and *-* None select the harmonic force constants without a cutoff radius.
Run alm to obtain the displacement patterns
$ ${ALAMODE_ROOT}/_build/alm/alm ALM0.in > ALM0.log
This finishes quickly and writes the two displacement patterns to pbte444.pattern_HARMONIC.
Then, create the displaced structures with displace.py
$ python ${ALAMODE_ROOT}/tools/displace.py --VASP POSCAR --mag 0.01 --prefix harm -pf pbte444.pattern_HARMONIC
Note that the primitive POSCAR is given, not a supercell.
displace.py reads the supercell built by alm from the header of the pattern file, so it needs PyYAML.
It generates two displaced supercells (harm1.POSCAR and harm2.POSCAR) with a displacement of 0.01 Å,
together with the supercell without displacements (SPOSCAR0), which is used as the reference structure in the next step.
2. Run DFT calculations and get forces#
Next, please perform DFT calculation and compute forces for the two displaced configurations.
The INCAR and KPOINTS.super (a \(2\times 2\times 2\) \(\Gamma\)-centered mesh for the supercell) are in vasp_inputs/.
The calculations can be done, for example, as
#!/bin/bash

cp vasp_inputs/INCAR INCAR
cp vasp_inputs/KPOINTS.super KPOINTS
for ((i=1;i<=2;i++))
do
    cp harm${i}.POSCAR POSCAR
    mpirun vasp_std
    mv OUTCAR OUTCAR.${i}
    mv vasprun.xml vasprun_${i}.xml
done
cp vasp_inputs/POSCAR POSCAR
Here, the last line restores the primitive POSCAR, which is used by alm in the next step.
When all calculations are done, please collect the displacement-force datasets as
$ python ${ALAMODE_ROOT}/tools/extract.py --VASP SPOSCAR0 vasprun_?.xml > DFSET_harmonic
The reference structure must be SPOSCAR0, not the primitive POSCAR:
extract.py computes the displacements by comparing the atomic positions in vasprun_?.xml with those in the reference structure atom by atom,
so the reference must be the undisplaced supercell with the same atom order as the supercell built by alm.
In this tutorial, we have placed the VASP outputs in vasp_outputs/ directory, so you can skip the VASP calculations by using the vasprun_?.xml files in that directory as
$ python ${ALAMODE_ROOT}/tools/extract.py --VASP SPOSCAR0 vasp_outputs/vasprun_?.xml > DFSET_harmonic
reference/SPOSCAR0 and reference/DFSET_harmonic are also provided.
3. Fit second-order force constants#
The input for the fitting (ALM1.in) is the same as ALM0.in except for MODE = optimize and the &optimize field.
&general
 PREFIX = pbte444
 MODE = optimize
 STRUCTURE_FILE = POSCAR
 SUPERCELL = 4
/

&interaction
 NORDER = 1
/

&cutoff
 *-* None
/

&optimize
 DFSET = DFSET_harmonic
/
Please run
$ ${ALAMODE_ROOT}/_build/alm/alm ALM1.in > ALM1.log
and check if the fitting error in the log file (ALM1.log) is reasonably small (1.24% in reference/ALM1.log).
The second-order force constants are saved in pbte444.h5, which also stores the crystal structure and is used by anphon below.
A pre-generated pbte444.h5 is provided in reference/.
4. Compute dielectric tensor and Born effective charges#
To include the non-analytic correction, one needs to give the electronic permittivity tensor (\(\epsilon_{\infty}\)) and the Born effective charges. This can be done conveniently using density functional perturbation theory (DFPT), which is available in major DFT codes (e.g., VASP, Quantum-ESPRESSO, abinit).
In VASP, for example, you just need to add LEPSILON = .True. in INCAR. For orbital dependent functionals, this option is not supported, and LCALCEPS = .True. should be used instead.
The inputs used for this tutorial are INCAR.dfpt and KPOINTS.dfpt (a \(20\times 20\times 20\) \(\Gamma\)-centered mesh) in vasp_inputs/, together with the primitive POSCAR.
System = PbTe_dfpt
ENCUT = 600
PREC = Accurate
ADDGRID = .True.
LREAL = .False.
GGA = PS
ISMEAR = 0; SIGMA = 0.01
EDIFF = 1.0E-9
LEPSILON = .True.
IBRION = -1; NSW = 0
LWAVE = .False.; LCHARG = .False.
Important
Please run a DFPT calculation using the same primitive cell (POSCAR) that you gave to alm to generate the supercell. If these two structures are inconsistent, the order of the atoms and Born effective charges may change, which likely results in strange phonon band dispersion.
Once the DFPT calculation finishes, please parse the electronic permittivity tensor and Born effective charges from the output file and save them in a single file following the specific format described in Format of BORNINFO.
If you used VASP, the BORNINFO file can be created by issuing
$ python ${ALAMODE_ROOT}/tools/extract.py --VASP POSCAR --get born vasprun.xml > PbTe.born
Here, POSCAR is the primitive cell used in the DFPT calculation, and vasprun.xml is the output of the DFPT calculation.
This parsing feature is supported only for VASP and Quantum-ESPRESSO.
We provide the pre-generated file PbTe.born in the reference/ directory, so you can copy it to the working directory to move forward.
It gives \(\epsilon_{\infty} = 30.40\) and \(Z^{*}_{\mathrm{Pb}} = -Z^{*}_{\mathrm{Te}} = 5.88\).
The Born effective charge changes by about 0.6% from a \(12\times 12\times 12\) to the \(20\times 20\times 20\) mesh, so please check the \(k\)-point convergence for your own system.
5. Phonon calculations#
All inputs are ready, so we can start phonon calculations.
We first compute phonon dispersion without non-analytic (NA) correction.
The input file for anphon (phband.in) looks like
&general
 PREFIX = PbTe_NA0
 MODE = phonons
 FCSFILE = pbte444.h5
 NONANALYTIC = 0; BORNINFO = PbTe.born
/

&kpoint
1
G  0.00000  0.00000  0.00000  X  0.50000  0.50000  0.00000 41
X  0.50000  0.50000  1.00000  G  0.00000  0.00000  0.00000 41
G  0.00000  0.00000  0.00000  L  0.50000  0.50000  0.50000 41
/
The crystal structure is read from pbte444.h5, so &cell and KD are not needed.
Copy it to the working directory (if you skipped the previous steps, copy reference/pbte444.h5 and reference/PbTe.born as well).
Please run anphon and plot the bands file
$ cp reference/phband.in .
$ ${ALAMODE_ROOT}/_build/anphon/anphon phband.in > phband.log
$ python ${ALAMODE_ROOT}/tools/plotband.py PbTe_NA0.bands
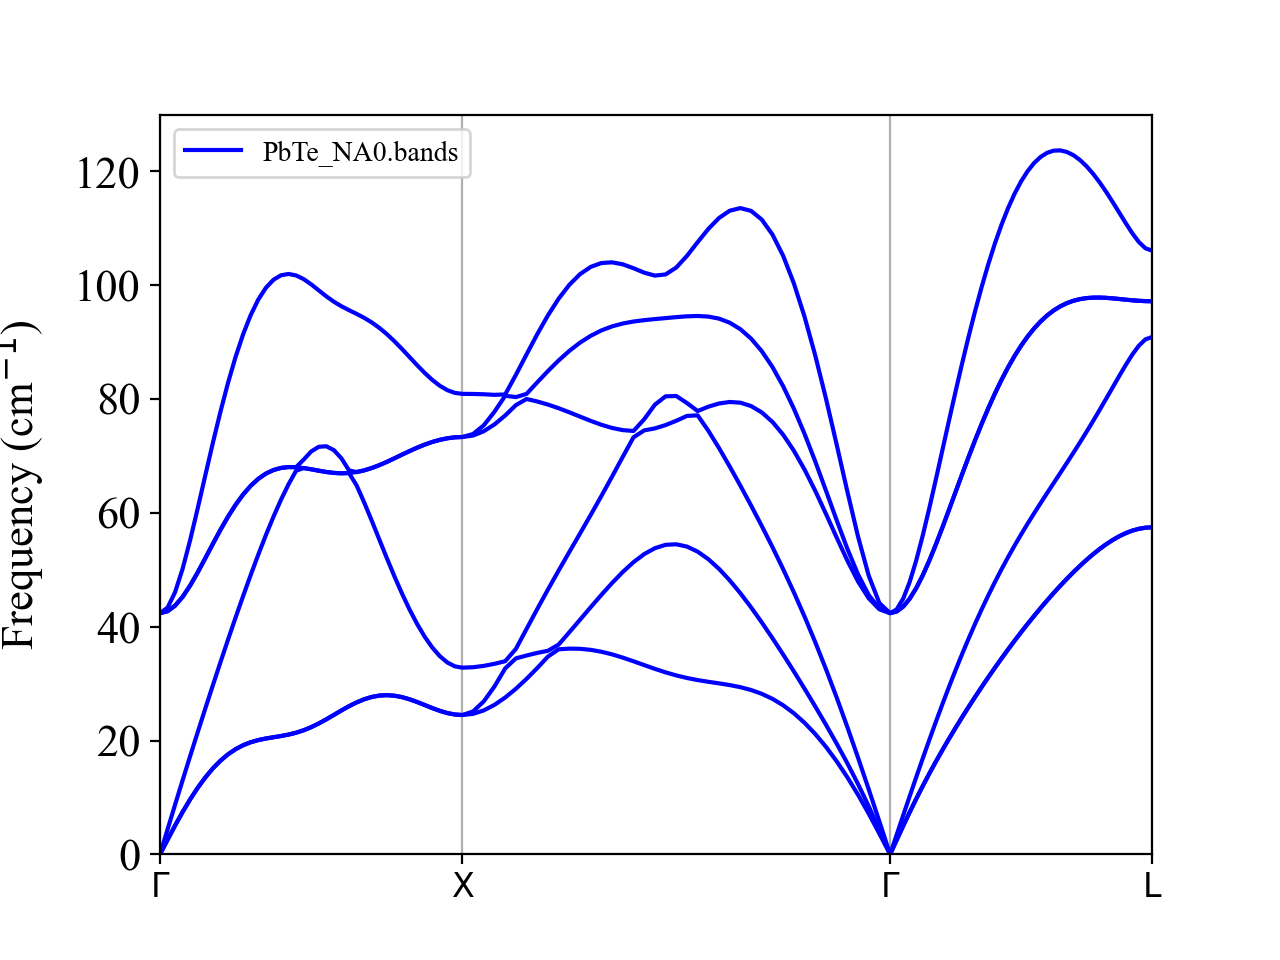
We can see that the TO and LO phonons are degenerate at \(\boldsymbol{q}=0\) and the optical phonon frequencies go down rapidly as \(\boldsymbol{q}\longrightarrow 0\). All three optical modes are at 42.4 cm-1 at \(\Gamma\).
Next, let us consider the NA correction.
Please change the PREFIX and NONANALYTIC tags as
PREFIX = PbTe_NA1
NONANALYTIC = 1; NA_SIGMA = 0.15
This option activates Parlinski’s approach with the damping factor of \(\sigma = 0.15\). After running anphon again, we can compare phonon dispersion curves with and without the NA correction.
$ ${ALAMODE_ROOT}/_build/anphon/anphon phband.in > phband.log2
$ python ${ALAMODE_ROOT}/tools/plotband.py PbTe_NA0.bands PbTe_NA1.bands
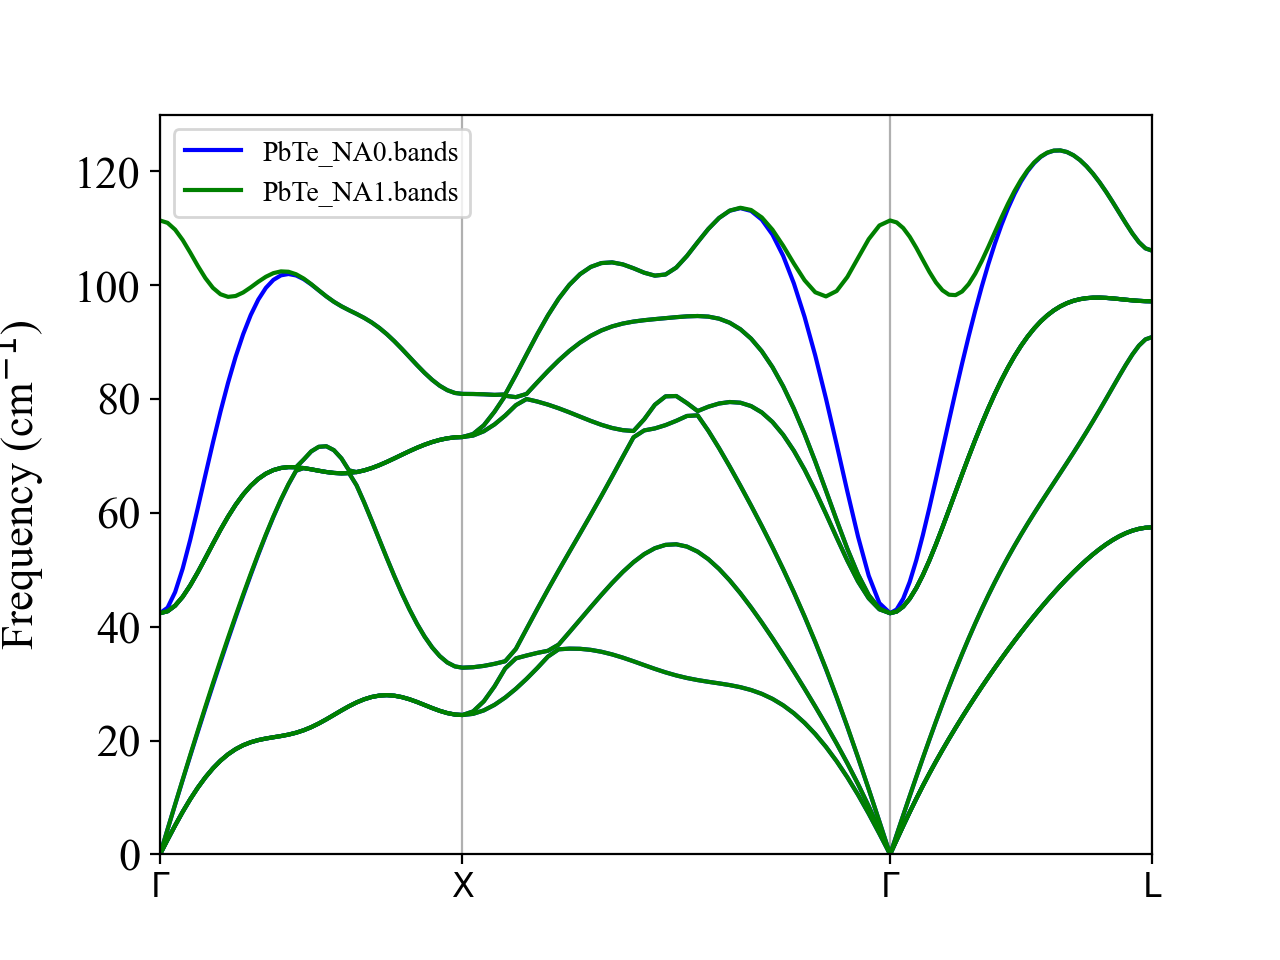
The LO-TO splitting occurs as expected: the LO frequency at \(\Gamma\) is now 111.3 cm-1, while the TO frequency stays at 42.4 cm-1.
Please repeat the above calculations with the other NONANALYTIC options (NONANALYTIC=2 (mixed-space) and NONANALYTIC=3 (Ewald method)).
Finally, we plot all results together to see the difference.
$ python ${ALAMODE_ROOT}/tools/plotband.py PbTe_NA?.bands
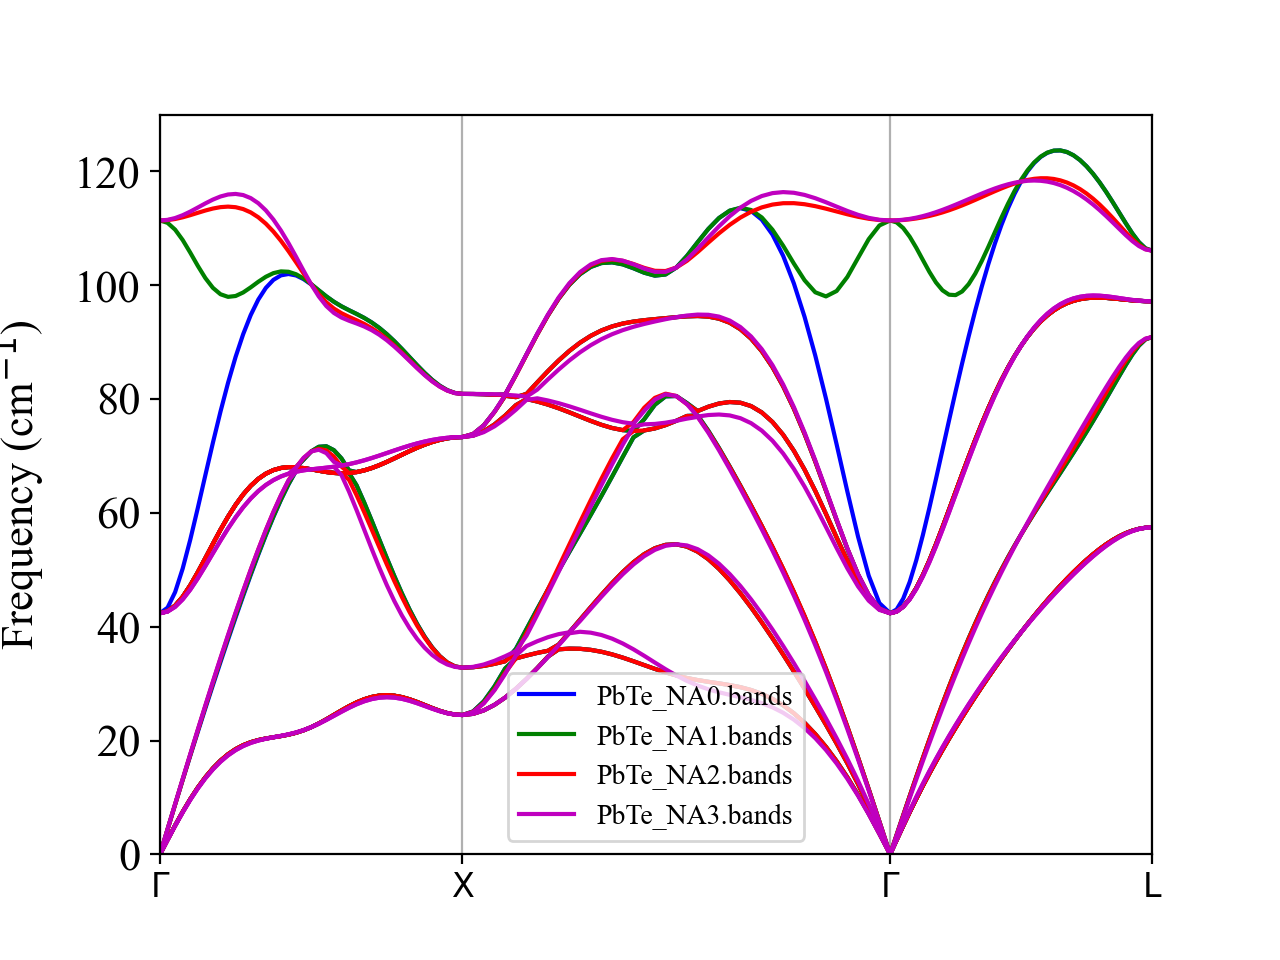
We can see that NONANALYTIC = 2 and NONANALYTIC = 3 give smoother dispersion than NONANALYTIC = 1.
For these two methods,the NA correction becomes exactly zero at the commensurate \(\boldsymbol{q}\) points (other than \(\boldsymbol{q}\longrightarrow 0\)). In the present calculation, we used a 4x4x4 supercell of the primitive cell. Hence, the NA correction should be zero at the Brillouin zone boundaries (X and L points) as well as the midpoint of each high-symmetry line (e.g., \(\boldsymbol{q}=(1/4, 1/4, 0)\)). We can confirm this from the above figure, where the blue, red, and magenta curves perfectly intersect with each other at these commensurate points. The NONANALYTIC = 1 result (green) instead shows unphysical dips of the LO branch between \(\Gamma\) and the zone boundaries, which are an artifact of this approach.
In general, we recommend using NONANALYTIC = 3 as it correctly separates the short- and long-range terms and calculates the NA correction in the reciprocal space accurately using the Ewald summation method. It can also avoid abrupt jumps or discontinuities in dispersion curves at Brillouin zone boundaries, which can happen with NONANALYTIC = 2 when the BZ boundary is not commensurate with the supercell size.